 EN
Home Skincare
Ideal for daily facial care routines, providing easy-to-use beauty solutions at home.

2026/10/07
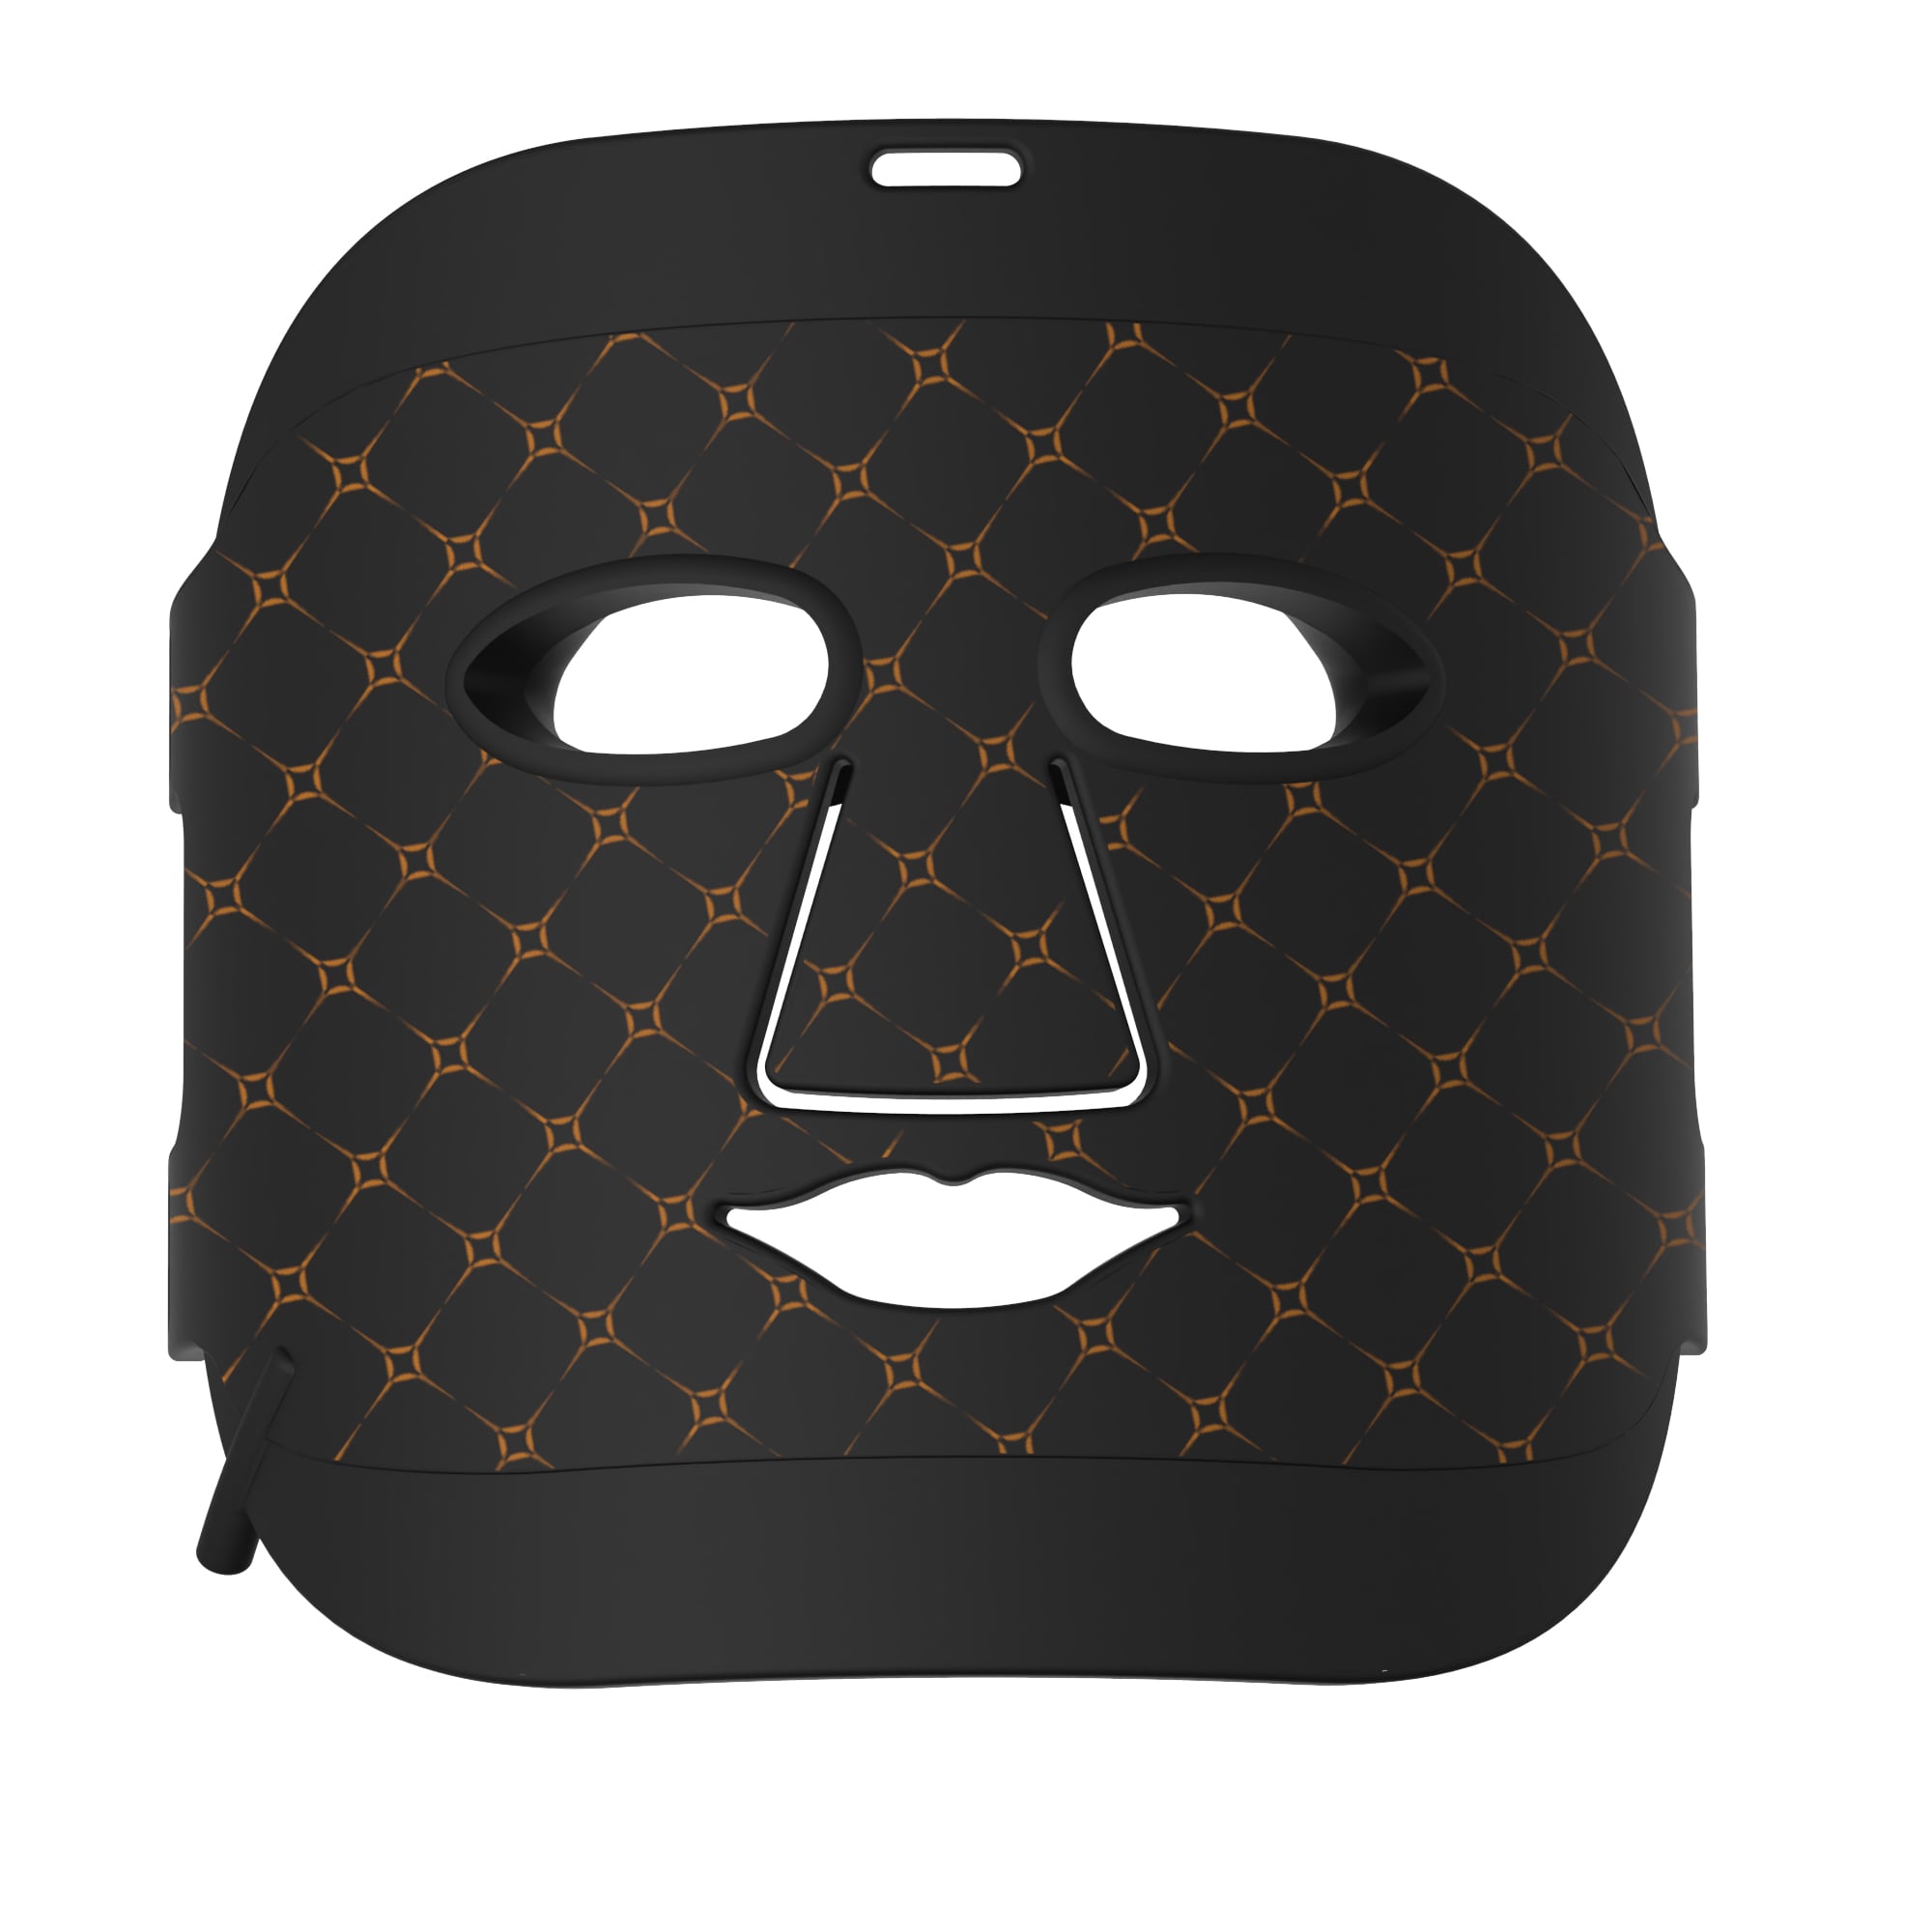
For a brand preparing to sell a led light therapy mask in the United States, the first compliance question is not whether the factory is “FDA registered.” It is what the product is intended to do. That distinction decides whether a led light therapy mask is regulated as a medical device, what classification may apply, and whether a premarket submission is needed before commercial distribution.
From an international buyer’s perspective, this is where procurement errors often start. A supplier may show a registration record, a test report, or a familiar-looking mask, yet none of those items by itself proves that a particular led light therapy mask is cleared for the claims on its packaging. The practical goal is to match the device, its indication, its evidence, and its labeling before launch.
FDA regulation turns on intended use and claims. A led light therapy mask marketed only with general cosmetic language may be assessed differently from one promoted to treat wrinkles, mild-to-moderate acne, or another disease-related condition. The same hardware can create a different regulatory question when the wording, instructions, target user, or stated result changes.
That is why a buyer should freeze the proposed claims early. Ask what the led light therapy mask will say on the product page, carton, instructions for use, social posts, and retailer copy. “Improves appearance” is not automatically a safe shortcut, and a medical-looking claim cannot be supported by a cosmetic label. Have a qualified regulatory professional review the exact U.S. claims for the finished product.
FDA says that establishment registration and device listing do not denote FDA approval, clearance, or authorization. In other words, an FDA registration number for a factory or a listed device is not proof that a led light therapy mask has passed 510(k) review. FDA also does not “certify” devices, so “FDA certified” is language to avoid.
A clearer way to assess a proposed led light therapy mask is to request the specific premarket reference, if one exists, then compare the manufacturer, model, indication, product code, and cleared labeling. A legitimate 510(k) record is not a blanket passport for a product family. It supports a particular device and intended use described in that submission.
Many light-based OTC wrinkle-reduction devices are regulated as Class II devices, but classification must be verified for the actual intended use. FDA materials for one cleared home-use LED mask, K250966, describe an OTC wearable light-based device for treatment of full-face wrinkles under 21 CFR 878.4810 and product code OHS. That example is useful because it shows that a led light therapy mask clearance is specific: it does not establish clearance for every color mode, claim, model change, or acne indication.
For a traditional 510(k), the submitter generally demonstrates that the new led light therapy mask is substantially equivalent to a legally marketed predicate device. The comparison begins with the same intended use. It also addresses technological characteristics and whether differences raise new questions of safety or effectiveness.
In plain terms, the submission should give FDA enough information to understand the led light therapy mask, its user population, operating mode, wavelengths, exposure parameters, controls, materials that contact the user, labeling, and comparison to the predicate. The exact evidence depends on the device and claim. A team should not assume that a competitor’s 510(k) can simply be copied or that clinical data is required in every case; FDA evaluates the full submission and the evidence needed for that device.
A reliable development file for a led light therapy mask normally links the design to its risk controls and proposed claims. Depending on the product, that can include performance verification, electrical safety and electromagnetic compatibility work, biocompatibility support for skin-contacting components, software or usability documentation where relevant, and labeling that explains safe operation. The final plan should come from the applicable FDA requirements and a device-specific regulatory assessment.
Do not treat a component report as proof for a finished led light therapy mask. Changes in LED layout, wavelength range, dose, treatment time, eye protection, controller, strap design, charger, materials, or intended population can affect the comparison and testing rationale. This is the point at which a disciplined supplier review saves time: map every marketed specification to controlled design documentation, not to an unverified sales sheet.
Suppose a led light therapy mask was cleared for full-face wrinkles. Using that history to advertise treatment of acne, rosacea, wound healing, pain, or a different patient group is not a minor marketing edit. It may change the intended use and require a new regulatory analysis. The same caution applies to “medical grade,” before-and-after claims, and implied disease claims in influencer content.
Here is a useful self-correction: it is tempting to ask whether a mask is “FDA cleared,” full stop. More accurately, ask whether this exact led light therapy mask is cleared, for this exact indication, under this manufacturer and model identity, with labeling that matches the marketed product. That question is more work, but it produces a defensible launch file.
FDA clearance is not a generic badge for an LED product category. For a led light therapy mask making medical-device claims, the requirements begin with intended use and continue through classification, the appropriate premarket pathway, evidence, labeling, registration/listing obligations, and postmarket controls. Buyers who verify those pieces together are in a far better position to choose a compliant led light therapy mask and keep marketing aligned with the product’s documented regulatory status.
This article is general educational information, not legal or regulatory advice. FDA requirements, product classifications, and submission expectations can change, so confirm the current requirements for the final device with FDA resources and qualified U.S. regulatory counsel.

2026/10/30
Discover why in-house mold development helps an RF beauty device manufacturer improve product consistency, iteration speed, tooling control, and production readiness.

2026/10/30
Learn how to verify an RF beauty device manufacturer's R&D and prototyping capabilities using practical engineering workflows and audit checklists.

2026/10/30
Learn what shapes MOQ when choosing an RF beauty device manufacturer and how to plan a lower-risk first order with balanced platform and customization.

2026/10/30
Use this RF beauty device manufacturer evaluation checklist to compare safety evidence, OEM and ODM capability, quality control, and launch readiness.

2026/10/30
See how technology makes beauty at home more guided, consistent, and informed—plus a practical checklist for choosing devices wisely.

2026/10/30
Explore how AI and smart devices shape the future of beauty with responsive routines, clear guidance, trustworthy design, and user-controlled care.

Ideal for daily facial care routines, providing easy-to-use beauty solutions at home.

Perfect for spas and salons’ frequent treatments, enhancing client experiences and results.

Supports dermatologists with advanced devices for professional skin therapy sessions.

Delivers innovative solutions for full-body massage and relaxation with home devices.

Features anti-aging technologies to help reduce wrinkles and stimulate skin vitality.

Specialized devices for safe and effective hair removal suitable for home or clinic use.

Our customers love the Perfect Idea beauty devices. Quality, packaging, and results are all outstanding. We saw increased repeat sales after adding their facial cleansers to our lineup. Their support team is responsive and reliable.

Perfect Idea brings innovation and reliability to home beauty appliances. Our collaboration introduced their massage tools to Japanese women, boosting customer satisfaction and growing our brand trust across the region.

We partnered with Shenzhen Perfect Idea Tech for their beauty equipment. The shipping was fast and every product matched our expectations. Their home-use massagers are especially popular with local salons and home users.

The stylish packaging and attractive design make Perfect Idea stand out. French consumers appreciate the easy instructions and visible results. Communication was efficient and orders always arrived on time.

Perfect Idea home use personal care devices have become a favorite with our clients. The beauty device line offers quality at a reasonable price, and technical training for our staff made onboarding easy.

We selected Perfect Idea for our beauty product portfolio and have seen very positive feedback from our customers. Products are easy to market online and the customer support is reliable and helpful.
